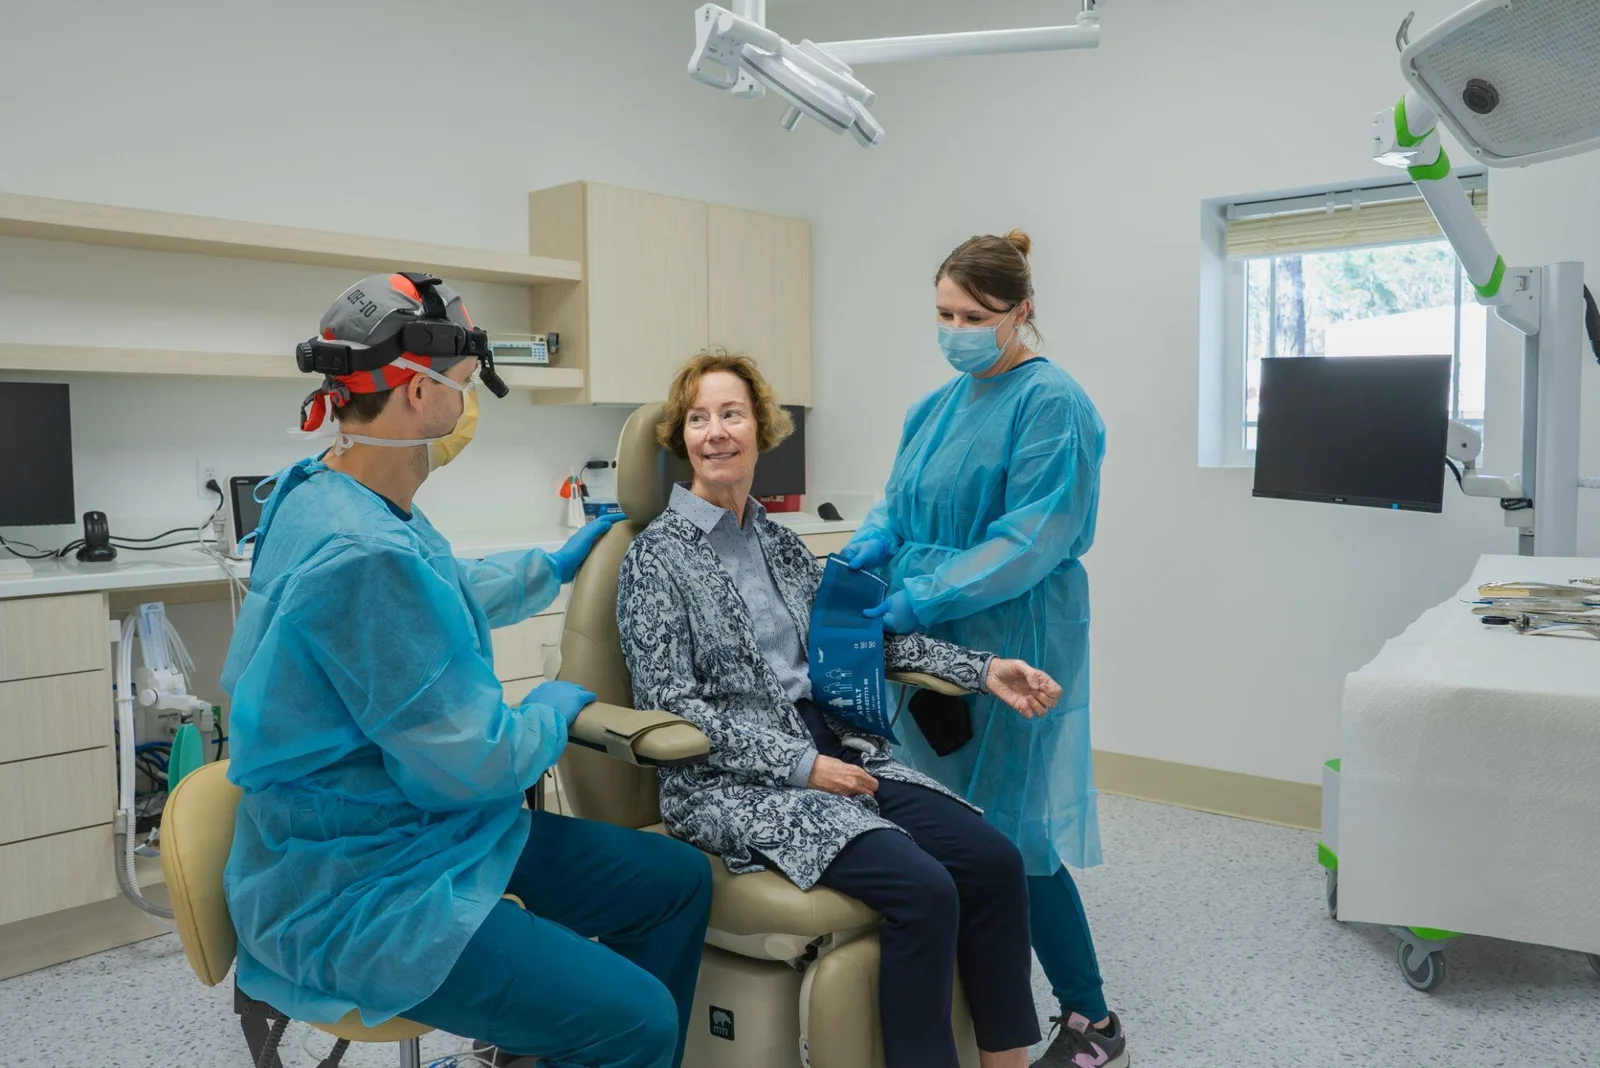

Photo: Dr. Dexter Mattox
For years, informed consent has largely functioned as a regulatory checkpoint in clinical research: present the paperwork, explain the protocol, document the agreement, and check the box.
That rigid framing is becoming increasingly difficult to justify as clinical trials undergo a rapid digital transformation. The fundamental question is no longer just whether a participant signed a consent form, but whether the process provided a genuinely meaningful opportunity to understand what they were agreeing to.
Challenging the Digital Assumption
Recent research suggests that format plays a far more decisive role than previously thought. In a randomized study published in npj Digital Public Health, researchers evaluated electronic consent against a traditional conversation-based process among 604 prospective biobank participants. The eConsent cohort achieved comprehension scores that were at least as strong as, and in several metrics superior to, the traditional paper-and-talk approach.
While these findings do not prove that digital consent is a universal remedy, they dismantle a long-held assumption: that shifting consent to a digital screen inherently dilutes informed decision-making.
This nuance is critical. Regulatory bodies like the U.S. Food and Drug Administration (FDA) have acknowledged that electronic consent involves much more than swapping paper for a digital signature. The FDA’s regulatory guidance explicitly addresses using electronic media to present trial details, permitting platforms to integrate text, graphics, audio, and interactive video.
However, the core obligation remains unchanged: participants must fully understand the scope, risks, and obligations of the study. Technology is not meant to replace human judgment or regulatory compliance, it is a vehicle to make complex, overwhelming clinical decisions easier to navigate.
The Double-Edged Sword of Digital Interfaces
When executed thoughtfully, a digital consent experience transforms how information is absorbed. It breaks dense protocols into manageable, digestible sections, lets participants revisit complex material at their own pace, embeds visual diagrams for difficult medical concepts, and uses quick self-assessment questions to verify understanding in real time. These features grant participants agency over their decision-making process in ways static paper documents never could.
Yet, going digital does not automatically make a process participant-friendly.
A poorly designed platform creates novel friction points. Participants possess varying degrees of digital literacy, face accessibility hurdles, or hit unexpected technical glitches. An overloaded interface that piles on interactive features without clarifying the underlying message can easily cause cognitive fatigue. A scoping review of decentralized clinical trials (DCTs) emphasized that digital literacy, user interface design, and real-time technical support require continuous attention as research moves offsite.
The takeaway for trial sponsors and principal investigators is clear: participant experience must be engineered before the consent process ever begins.
The goal should never be shortening consent forms just because they are displayed on a tablet. The goal is making complex scientific information navigable, transparent, and appropriately interactive, all while preserving space for human discussion with the research team.
Bridging the Gap Between Design and Equity
Designing for participant experience also requires an intentional focus on health equity and diverse patient populations. In traditional paper trials, complex medical jargon and dense legal disclaimers often create disproportionate barriers for underrepresented communities, non-native language speakers, or individuals with low health literacy. Digital platforms offer the technical capability to break down these barriers, providing real-time translation options, adjustable font sizes, screen-reader compatibility, and interactive glossaries that define complex terms on demand. However, if interface designers fail to account for cultural nuance or varying access to high-speed internet, digital consent risks exacerbating the very health disparities it has the potential to solve.
To prevent eConsent from becoming an unintended barrier, trial sponsors are increasingly adopting co-design methodologies, testing consent interfaces with actual patient advisory boards and advocacy groups before launching a study. By gathering feedback on visual clarity, navigation speed, and content readability, research teams can ensure that the technology accommodates real-world user behavior. Ultimately, a digital consent workflow should feel as accessible and intuitive to an elderly participant on a low-bandwidth cellular connection as it does to a tech-savvy participant using the latest tablet. Equity in clinical trials begins with making the entry point equally navigable for everyone.
From One-Time Compliance to Ongoing Engagement
Consent also serves as the foundation for the entire participant-investigator relationship. A participant who thoroughly grasps a study’s schedule, procedures, time commitments, and potential side effects enters the trial with calibrated expectations. While transparent eConsent does not guarantee lower drop-out rates on its own, it directly influences how participants perceive and trust the study from day one.
Industry leaders like AXIS Clinicals have advocated for this broader operational perspective, framing digital consent not as a static compliance event, but as an evolving, participant-centered interaction.
Crucially, eConsent cannot operate as an isolated tool. It must fit seamlessly into the broader clinical trial infrastructure, interoperating cleanly with Electronic Data Capture (EDC), eSource platforms, version control systems, and audit trails.
This system integration is vital because participant experience does not terminate once a digital signature is captured. Throughout a trial, participants may need to review study guidelines, receive protocol updates, sign re-consent amendments, or submit patient-reported outcomes. If the consent interface feels disconnected or difficult to navigate, it introduces friction into every subsequent stage of the study.
Ultimately, the shift taking place across clinical research is subtle but profound. Informed consent will always fulfill an essential regulatory mandate, but its real-world function extends far beyond legal documentation. In an increasingly digital landscape, asking whether a participant signed their name is no longer enough.
The true measure of success is whether the experience helped that human being truly understand what they were choosing to do.